Epigenetics and methylation analysis
Comprehensive genomic and epigenomic analysis in one go
New benchmarking data demonstrates why Oxford Nanopore technology is the new gold standard for methylation sequencing.
Epigenetic modifications regulate gene expression and play crucial roles in development and disease. Oxford Nanopore sequencing allows you to streamline your epigenetics workflow by directly detecting DNA or RNA modifications at single-nucleotide resolution — including 5mC, 5hmC, 6mA, and m6A — alongside the nucleotide sequence. No additional sample preparation or sequencing runs are required.
Any-length reads further enable structural variants (SVs), single nucleotide variants (SNVs), and repeats to be detected and phased along with epigenetic modifications — providing comprehensive genomic and epigenomic analysis in one go.
Why nanopore sequencing?
Gold-standard methylation sequencing
Get best-in-class 5mC detection directly from native DNA, with no bisulfite conversion or additional sequencing runs.
Comprehensive analysis, one simple workflow
Detect and phase SNVs, SVs, repeats, and methylation in a single streamlined assay.
More complete CpG coverage
Characterise methylation at CpG sites inaccessible to conventional approaches.
Reveal RNA modifications directly
Gain isoform-level insights across full-length transcripts.
Flexible and scalable
Scale genome- or transcriptome-wide modification analysis from individual samples to population studies, or target genomic regions with Adaptive Sampling.
Discover the new gold standard for methylation sequencing
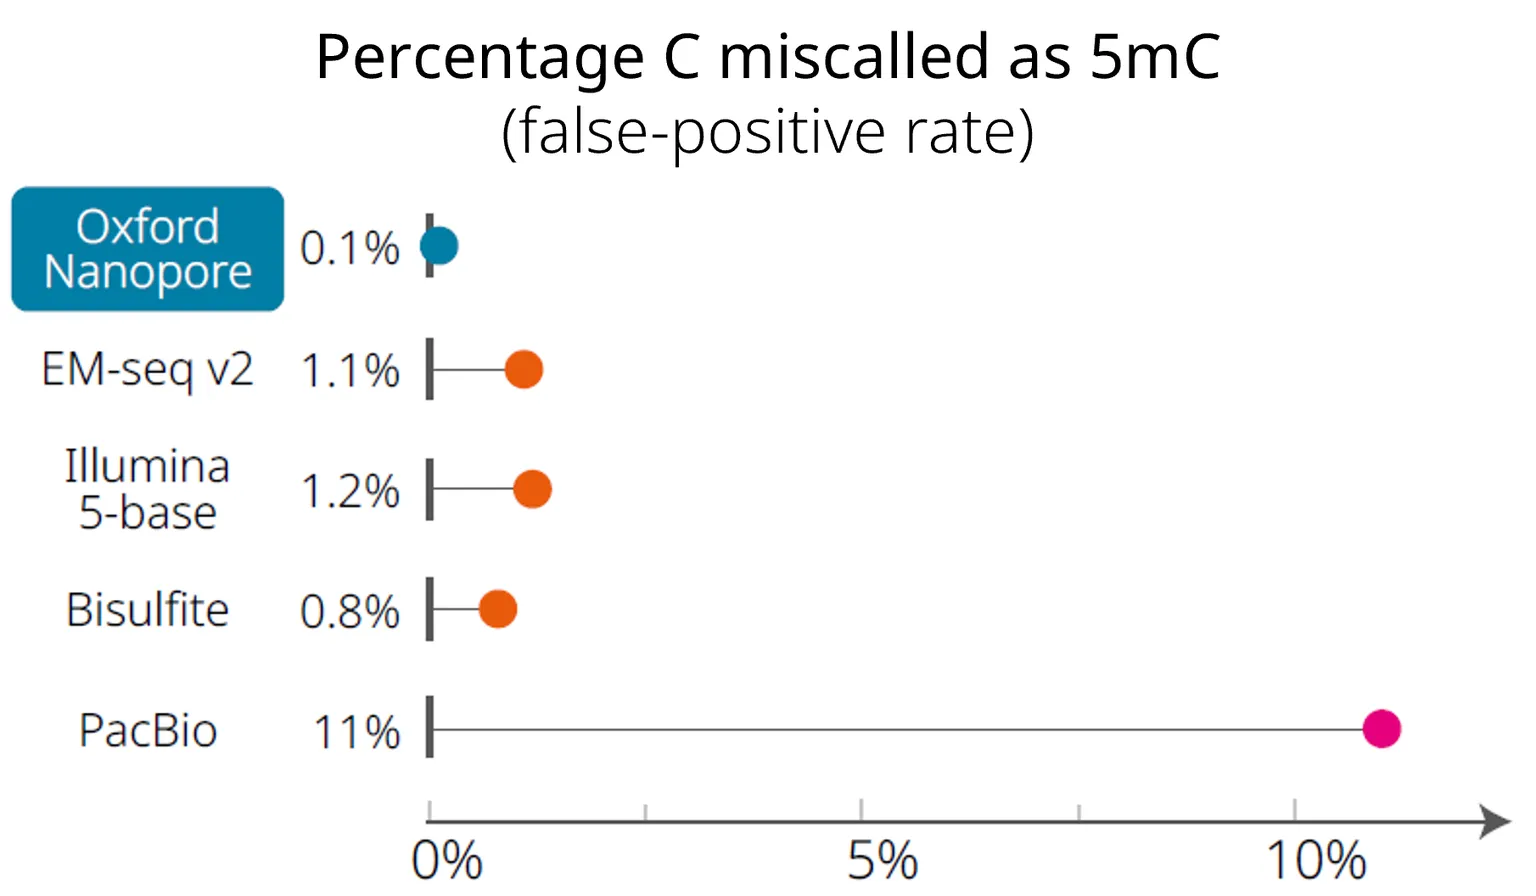
Oxford Nanopore sequencing delivers best-in-class 5mC calling accuracy with a false-positive rate of just 0.1% — lower than PacBio and all Illumina short-read methods benchmarked.
Compare benchmarking performance across false-positive and false-negative rates, CpG coverage, and more.
Customer spotlight
Population-scale variant and methylation analysis in a single assay
‘We concluded that the methylation data is really high quality’
Hear how Fritz Sedlazeck and the All of Us team profiled 27 million CpG sites per genome across 1,490 Hispanic individuals, while characterising small and structural variants and tandem repeats, with phasing from the same assay.
End-to-end workflows
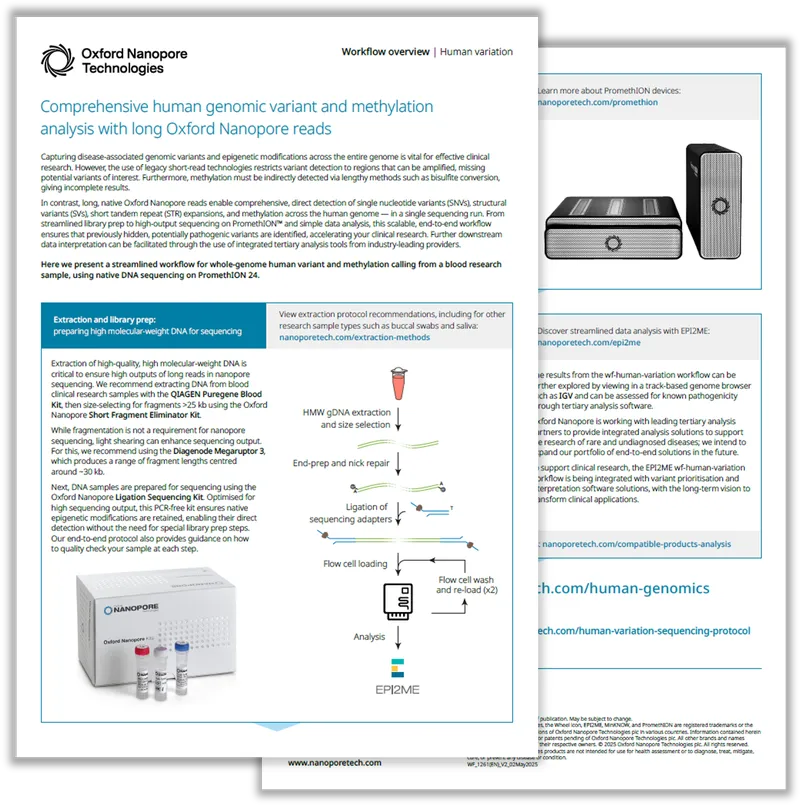
Comprehensive human genomic variant and methylation analysis with long Oxford Nanopore reads
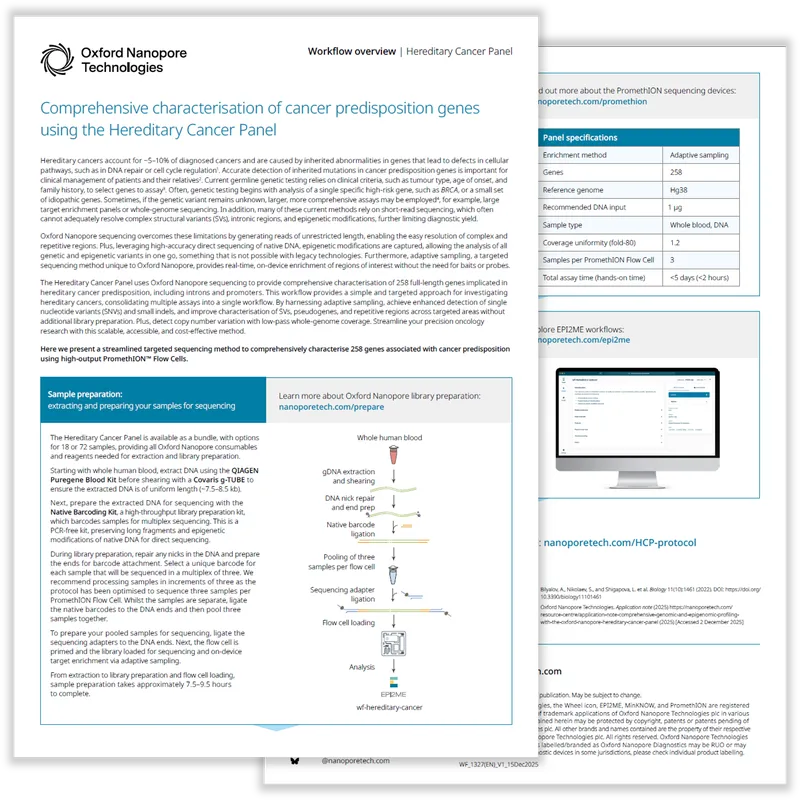
Comprehensive characterisation of cancer predisposition genes using the Hereditary Cancer Panel
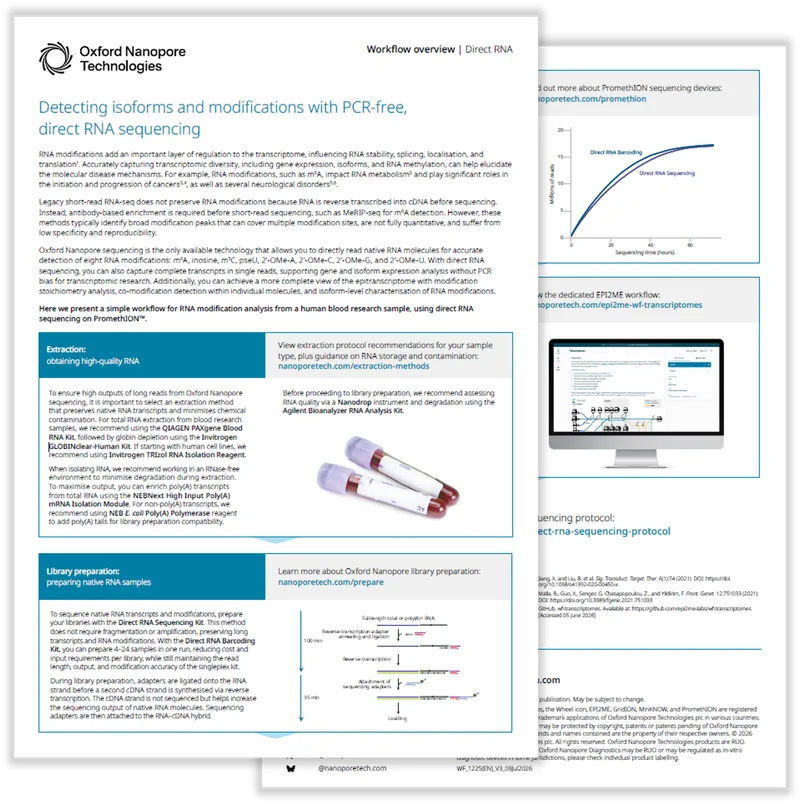
Detecting isoforms and modifications with PCR-free, direct RNA nanopore sequencing
Characterising genomic and epigenomic variation between tumour-normal research samples using long nanopore sequencing reads
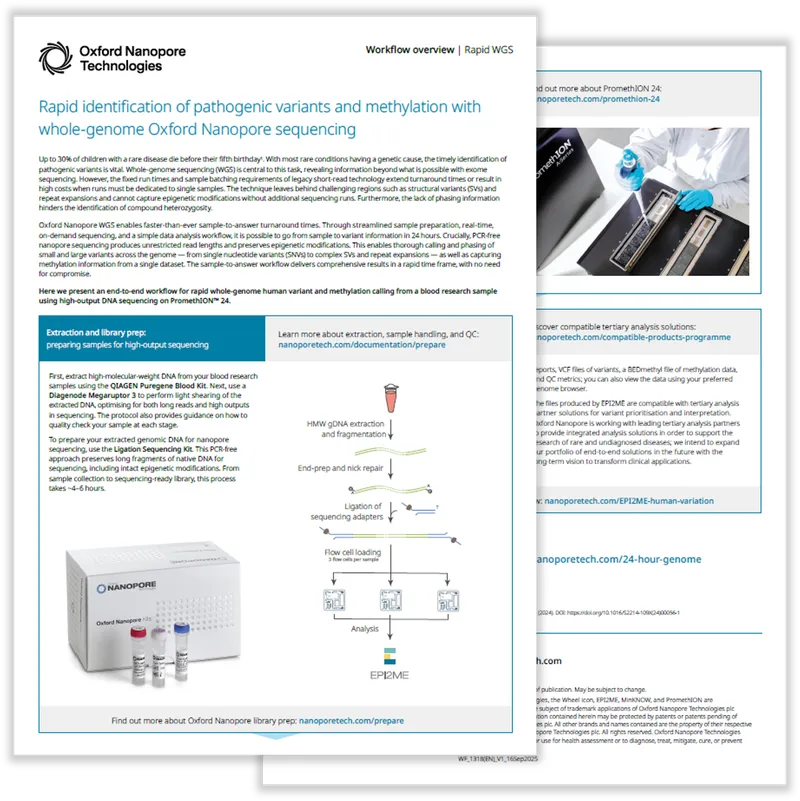
Rapid identification of pathogenic variants and methylation with whole-genome Oxford Nanopore sequencing
Featured resources
A guide to investigating methylation in the human genome
Oxford Nanopore technology allows you to detect DNA methylation alongside the nucleotide sequence, without additional library preparation steps. This guide provides a complete introduction to methylation sequencing in human genomes.
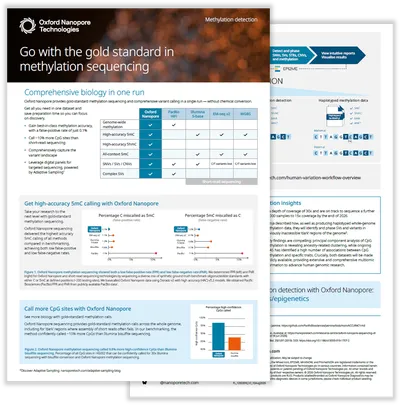

Demo: how to analyse methylation and variants in one go
Recommended device for epigenetics and methylation analysis

PromethION 24
Scale gold-standard methylation analysis across many samples. With up to 24 independently addressable flow cells and powerful onboard compute, PromethION 24 delivers flexible, on-demand access to terabases of sequencing data for comprehensive genomic, transcriptomic, and epigenomic analysis.
Streamlined analysis solutions
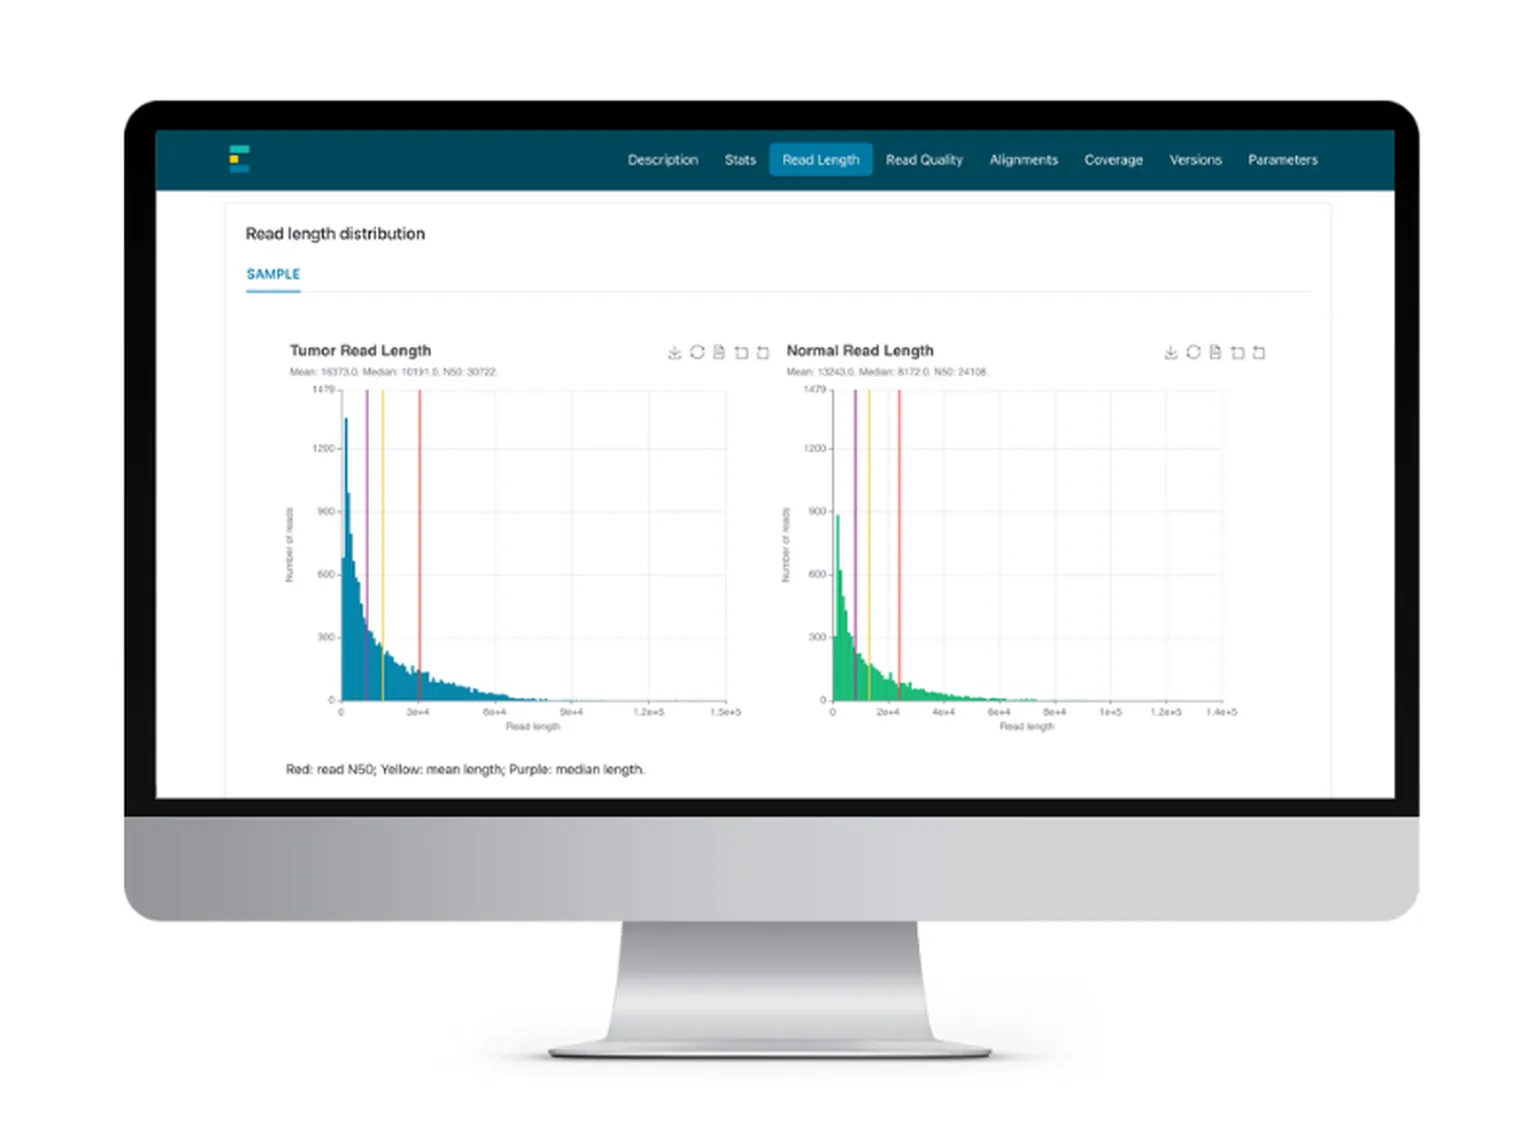
Get maximum insights from your data
Discover our comprehensive range of analysis tools — from intuitive, preconfigured workflows in EPI2ME to the latest analysis algorithms.
FAQs
What epigenetic modifications can Oxford Nanopore sequencing detect?
An expanding range of DNA and RNA modifications can be detected through the basecalling software Dorado, which is integrated into MinKNOW, the operating system for all Oxford Nanopore sequencing devices. Currently supported DNA modifications include 5mC, 5hmC, 6mA, and 4mC, while RNA modifications include m6A, pseudouridine (pseU), m5C, and inosine. Additional base modifications can be detected in the standalone version of Dorado.
Researchers have also developed tools for detecting further base modifications. Explore the latest publications in our Resource Centre.
How accurate is Oxford Nanopore methylation calling?
Internal benchmarking studies against PacBio and Illumina sequencing methodologies demonstrated that Oxford Nanopore sequencing delivers best-in-class 5mC calling accuracy, with exceptionally low false-positive and false-negative rates. View benchmarking data. View all sequencing accuracy metrics.
Does nanopore methylation sequencing require bisulfite conversion?
No. Oxford Nanopore sequencing detects base modifications directly from native DNA or RNA, so no bisulfite conversion or other chemical or enzymatic treatment is required. Base modifications are detected alongside the nucleotide sequence, without the requirement for additional sequencing runs.
Can Oxford Nanopore sequencing distinguish 5mC and 5hmC?
Yes. Oxford Nanopore sequencing can directly detect and distinguish 5mC and 5hmC — alongside other base modifications — from native DNA in a single assay. In contrast, conventional bisulfite sequencing does not distinguish between these modifications without additional approaches and sequencing runs.
Can methylation and genetic variants be detected in the same sequencing run?
Yes. Oxford Nanopore sequencing can call and phase base modifications alongside genetic variants such as SNVs, SVs, and repeats from the same sequencing data. Because native DNA (or RNA) is sequenced directly, base modifications are preserved and can be analysed together with the underlying nucleotide sequence, enabling genetic and epigenetic information to be generated in a single assay.
What software is used for Oxford Nanopore methylation calling?
Dorado performs high-accuracy canonical basecalling and modification calling, including methylation detection. Dorado is integrated into MinKNOW, the software used to operate Oxford Nanopore sequencing devices, and is also available as a standalone command-line tool.
For comprehensive human genome analysis, the EPI2ME workflow wf-human-variation provides methylation annotation and haplotype phasing alongside analysis of SNVs, CNVs, SVs, and STRs. It can be run through a point-and-click interface or from the command line.
Can I perform targeted sequencing without losing methylation information?
Yes, Adaptive Sampling enables PCR-free enrichment of regions of interest, allowing preservation and calling of base modifications. With no limit to the length of target regions, Adaptive Sampling is ideal for enrichment of very large regions from native DNA samples, such as megabase-scale SVs or whole chromosomes.


